DeepMind Launches AlphaGenome Atlas: An AI-Predicted Map of 9 Billion DNA Variants

AlphaGenome Atlas is the first AI-powered searchable database predicting functional effects of all 9 billion possible human single-base DNA variants.
AlphaGenome Atlas is an AI-powered database released by DeepMind that systematically predicts the biological functional impact of approximately 9 billion possible single-base substitutions across the human genome. With ~3 billion base positions each capable of three substitutions, traditional experiments cannot validate this vast variant space. The database packages AI model predictions into a directly queryable tool, with potential applications in variant functional screening from whole-genome sequencing, interpretation of variants of uncertain significance in clinical genetics, and non-coding regulatory region research. However, the database provides statistical predictions rather than experimental conclusions, and key details on model architecture, accuracy benchmarks, and access methods have yet to be fully disclosed — warranting particular caution in clinical settings.
What Is AlphaGenome Atlas
A searchable database covering the predicted effects of 9 billion single-base DNA variants has officially launched. According to its developers, AlphaGenome Atlas is an AI-powered resource designed to systematically map the predicted biological effects of every possible single-letter (base) change across the human genome.
A "single-base change" refers to replacing one of the four bases — A, T, C, or G — at a given position in the genome with another. The human genome contains approximately 3 billion base positions, and each can theoretically undergo three different substitutions, yielding roughly 9 billion possible combinations. The core value of AlphaGenome Atlas lies in providing systematic, AI-predicted annotations across this vast mutational landscape.
Why This Matters
Understanding the biological consequences of any given DNA mutation has long been a central challenge in genetics and basic biology research. The vast majority of variants fall within non-coding regulatory regions, where their functional effects are difficult to verify through traditional experiments — which are costly, time-consuming, and nowhere near capable of handling 9 billion combinations.
AI prediction models offer a fundamentally different approach. By learning from genomic sequences and functional data, these models can predict the functional impact of variants that have never been experimentally validated. AlphaGenome Atlas packages this predictive capability into a searchable database, meaning researchers can directly query predicted effects at sites of interest without needing to run complex models themselves.
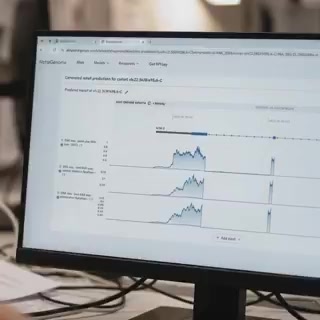
The core model behind AlphaGenome Atlas is AlphaGenome, released by DeepMind in 2024, which builds on earlier sequence-to-function prediction models such as Enformer. The fundamental idea behind these models is: take a segment of genomic DNA as input and predict a range of molecular phenotype outputs for that region — such as gene expression levels, chromatin accessibility (ATAC-seq signal), histone modification states, and transcription factor binding sites. By feeding in the reference sequence and an alternative sequence carrying the target variant separately, then comparing the predicted output differences, the model can quantify the predicted functional impact of that variant across various signals — a process known as in silico mutagenesis (ISM). Unlike earlier tools focused only on coding-region amino acid changes, these models can handle regulatory variants in non-coding regions. Since over 90% of variants associated with human genetic diseases fall in non-coding regions, this is a key advantage of the approach.
What This Means for Researchers
The developers emphasize that this tool aims to help researchers "better understand our biology." In practical terms, this type of resource has potential in several directions:
Accelerating Variant Functional Screening
When whole-genome sequencing uncovers a large number of variants, researchers often need to prioritize which are most likely to be functionally significant. A prediction map covering all possible variants can serve as an initial screening tool to narrow down candidates that warrant experimental validation.
Supporting Interpretation of Disease-Associated Variants
In clinical genetics and disease mechanism research, determining whether a rare variant is "pathogenic" or "benign" is a key challenge. AI-predicted functional impact scores can provide an additional reference dimension for such judgments, especially for novel variants with no prior reports.
In clinical genetics, variant interpretation typically follows the classification framework established by the American College of Medical Genetics and Genomics (ACMG), which categorizes variants into five tiers: "Pathogenic," "Likely Pathogenic," "Variant of Uncertain Significance (VUS)," "Likely Benign," and "Benign." Major variant databases worldwide (such as ClinVar) have accumulated millions of VUS records with unresolved clinical significance, posing major challenges for genetic counseling and patient management. AI functional prediction scores have the potential to serve as new ACMG evidence criteria — some tools like SpliceAI have already been recognized for splice site variants — potentially helping reclassify VUS entries. However, regulatory validation standards and clinical adoption thresholds remain significant barriers.
Providing Leads for Regulatory Region Research
The functional effects of non-coding variants have long been a blind spot in research, and sequence-based AI models have shown advantages in capturing regulatory signals. Systematic prediction data may reveal previously overlooked regulatory elements.
Non-coding regulatory regions encompass functional elements such as promoters, enhancers, silencers, insulators, and various non-coding RNA genes, which finely regulate gene expression across spatial and temporal dimensions. Traditional large-scale mutational effect assays — such as MPRA (Massively Parallel Reporter Assay) — can batch-test regulatory element activity in cellular models, but their throughput is still far from sufficient to cover 9 billion variants, and they struggle to recapitulate the true in vivo regulatory context across tissue-specific and developmental settings. Sequence-based AI models, trained on multi-omics functional data across dozens of cell types, capture cell-type-specific regulatory signals to some degree. This makes systematic prediction maps a promising "navigation tool" for designing targeted follow-up experiments — not a replacement for experiments themselves.
A Note of Caution
It's important to recognize that AlphaGenome Atlas provides "predictions," not "conclusions." The outputs of AI models are fundamentally statistical inferences based on existing data, and their accuracy is bounded by the coverage of training data and the model's inherent limitations. Researchers should still validate predictions experimentally, especially in contexts involving clinical decisions.
Based on currently available information, this launch primarily communicates product positioning and vision. Key technical details — including the model's specific architecture, benchmark accuracy evaluations, and database access methods — have yet to be fully disclosed. These details will directly determine the tool's practical value in real-world research settings.
Summary
AlphaGenome Atlas represents the deepening role of AI in basic biological research — moving from analytical assistance toward the systematic construction of queryable knowledge resources. Organizing the predicted effects of 9 billion DNA variants into a searchable database is itself a meaningful fusion of engineering and science. Whether it truly advances biological understanding will depend on the reliability of its predictions and the degree to which the research community adopts it. For researchers working at the intersection of computational biology and AI for Science, this is a development worth tracking closely.
Related articles

Reinforcement Learning Takes On Getting Over It: Distributed Training Strategies Behind 63% Progress
A developer uses deep reinforcement learning to tackle Getting Over It. The agent climbs from the bottom to 63% progress but gets stuck at the anvil. The project is open-sourced, using divide-and-conquer training and CPU-only training, seeking community collaboration to break the bottleneck.

Sigma FP2 Released: The World's Smallest Full-Frame Camera Gets Sensor and Screen Upgrades
Sigma launches the FP2 full-frame camera, retaining its smallest-and-lightest design with three key upgrades: improved sensor, articulating LCD screen, and longer battery life—built for portable video creators.

AAAI 2027 Phase 2 Decisions Imminent: Community Debates Review Fairness and Submission Ecosystem
AAAI 2027 phase 2 review results are imminent, with the Reddit community debating reviewer consistency and process transparency. This article examines pre-decision sentiment, structural tensions in top conference review, and practical strategies for authors.